💊 About About ADMETShiny
⚙️ Principal Functions of ADMETShiny in R
With ADMETShiny, say goodbye to manual ADMET analysis spread across multiple web servers. Manage all your physicochemical profiles in a single place.
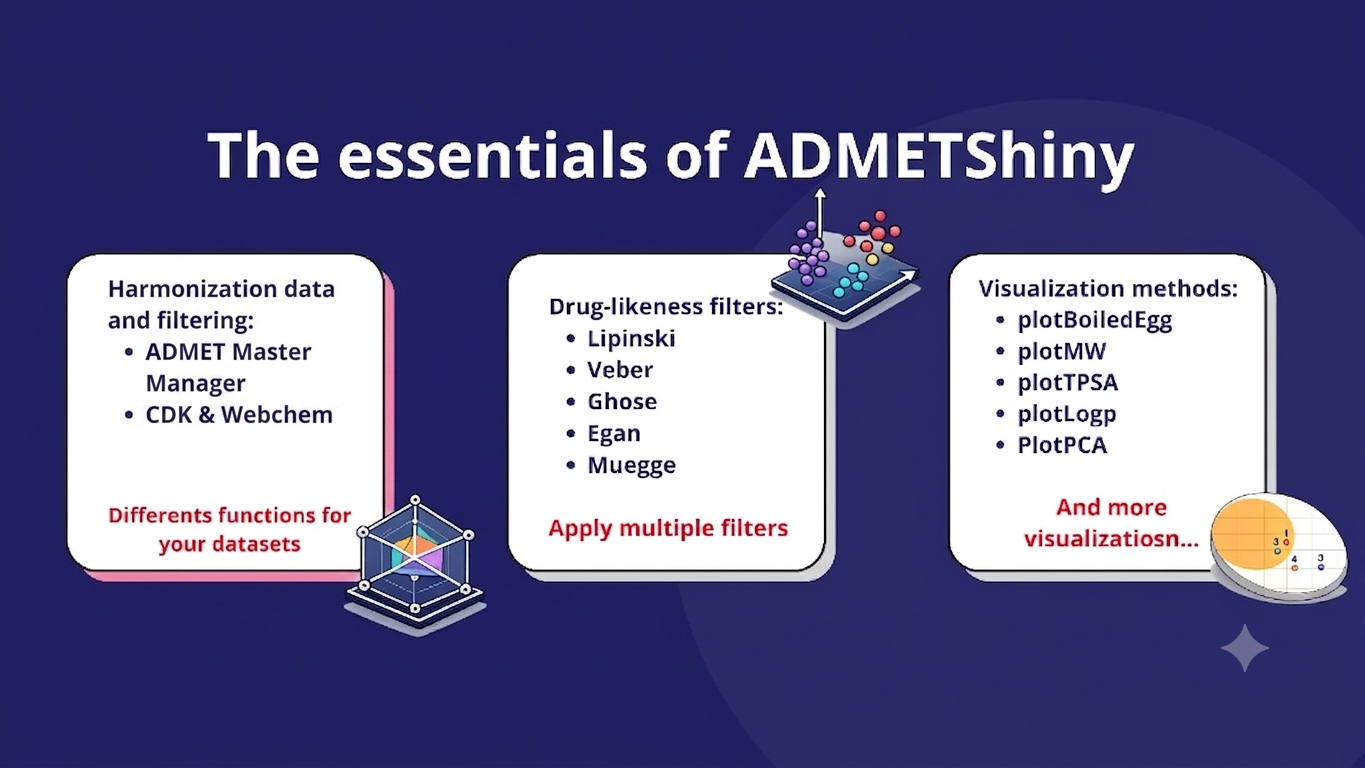
Once your analysis is complete, you can generate a single consolidated report summarizing the main characteristics of each model.
Don’t want to submit your compounds to external websites? Don’t worry. ADMETShiny includes a local web server built with the rCDK R package and the Java SE Development Kit (JDK), so your profiles can be estimated entirely on your own machine.
🌐 One for Open Source, and Open Source for One
ADMETShiny accepts input data from multiple sources:
| Input method | Details |
|---|---|
| 📄 Datasets | Upload files in .csv format |
| 🔎 PubChem search | By CID, common name, InChIKey, or CAS number |
🧪 Calculate drug-likeness properties with CDK and webchem
Dataset management and filters can be applied to the calculated molecular descriptors, producing clear and reproducible reports with minimal effort.
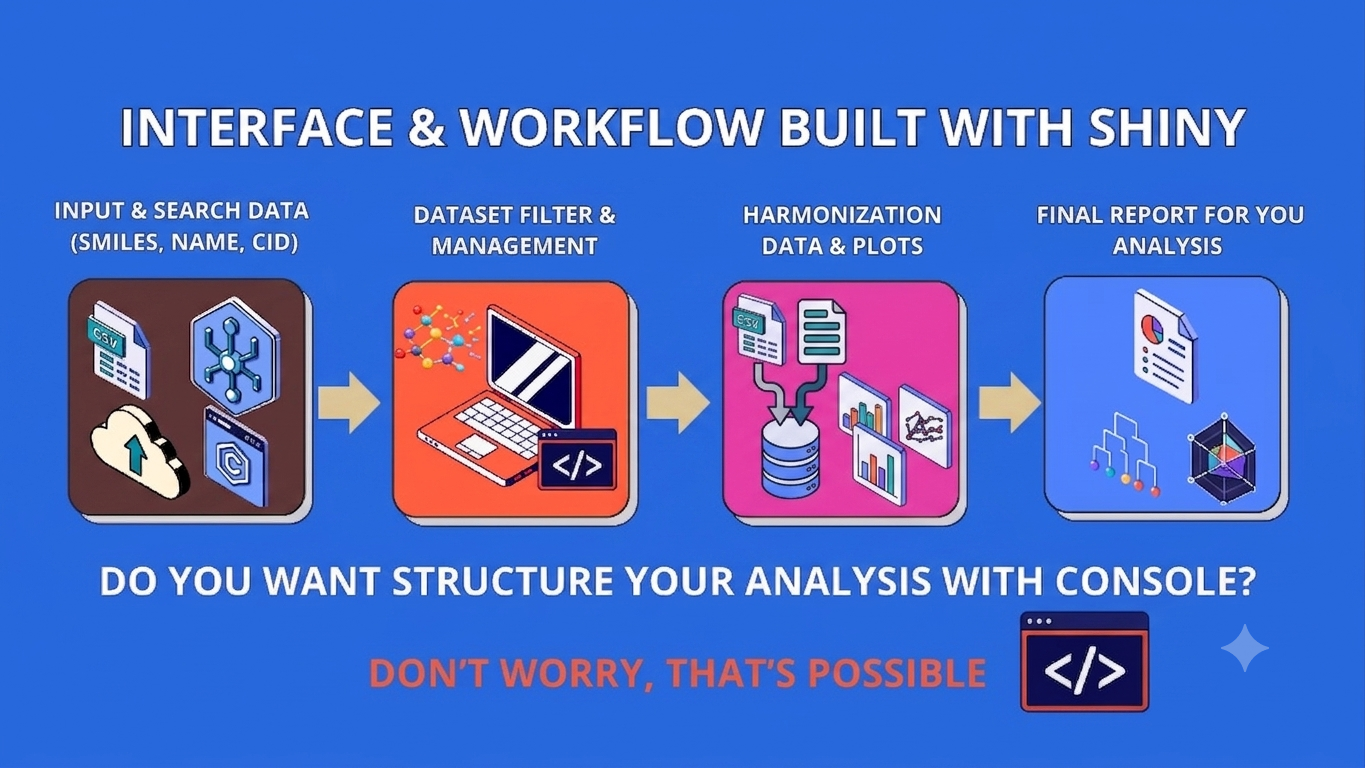
💻 Prefer the console? Build reproducible scripts
Everything available in the graphical interface can also be run from the R console, allowing you to develop simple, reproducible scripts and generate your reports programmatically.
👥 The Shiny App for Coders and Non-Coders
ADMETShiny offers an intuitive graphical interface that makes advanced cheminformatics analyses accessible without programming experience, while remaining fully integrated with the R ecosystem for users who require reproducible, customizable, and extensible workflows.
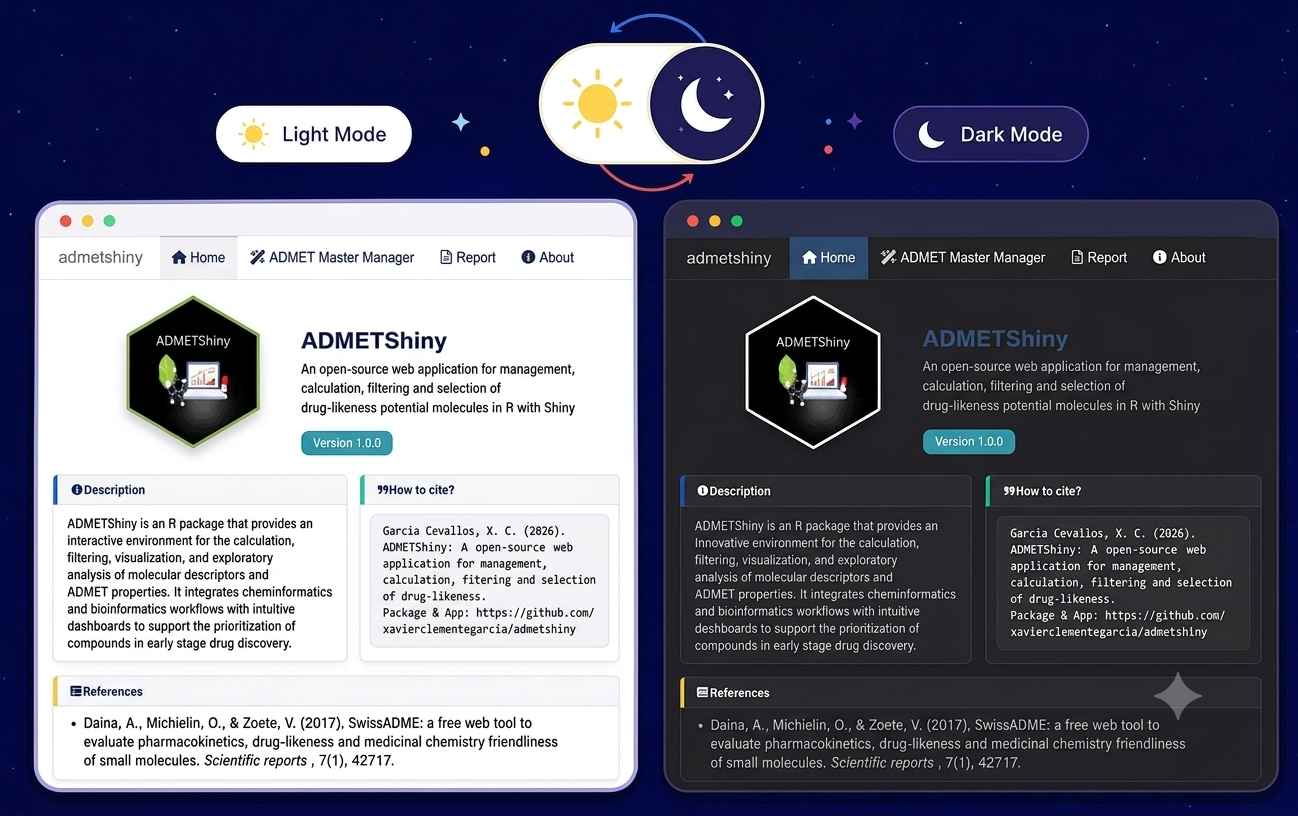
🧠 Knowledge Structure Analysis
ADMETShiny provides a comprehensive collection of interactive modules that streamline the early stages of drug discovery in a single, user-friendly environment:
- 🔬 Molecular retrieval
- 🧮 Descriptor calculation
- 🧬 ADMET prediction
- 🌌 Chemical space exploration
- 🔗 Structural similarity analysis
- 📑 Automated reporting
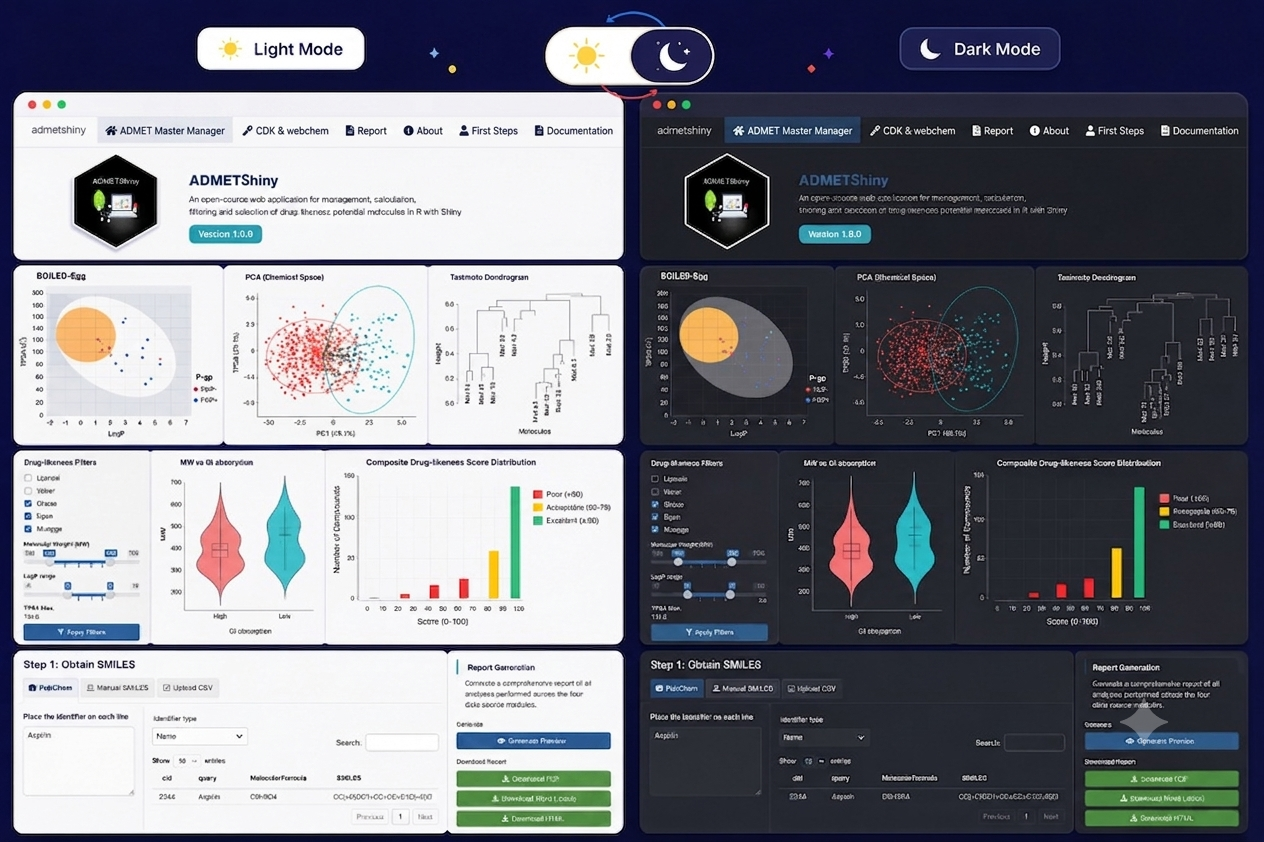
ADMETShiny combines data retrieval, descriptor calculation, predictive modeling, interactive visualization, molecular prioritization, and automated reporting in a unified open-source platform, designed to accelerate computational drug discovery and medicinal chemistry research.